Cut the phylogeny and categorical trait/range mapping for a given focal time in the past
Source:R/cut_simmaps_for_focal_time.R
cut_simmap_for_focal_time.RdCuts off all the branches of the phylogeny which are
younger than a specific time in the past (i.e. the focal_time).
Branches overlapping the focal_time are shortened to the focal_time.
Likewise, remove mapping for the cut off branches
by updating the $maps and $mapped.edge elements.
Arguments
- simmap
Object with the classes
"phylo"and"simmap", typically generated withphytools::make.simmap(), that contains a phylogenetic tree and associated categorical trait/range mapping. The phylogenetic tree must be rooted and fully resolved/dichotomous, but it does not need to be ultrametric (it can include fossils).- focal_time
Numeric. The time, in terms of time distance from the present, for which the tree and mapping must be cut. It must be smaller than the root age of the phylogeny.
- keep_tip_labels
Logical. Specify whether terminal branches with a single descendant tip must retain their initial
tip.label. Default isTRUE.
Value
The function returns the cut/updated simmap as a list of at least six elements with the classes "phylo" and "simmap":
Initial updated elements:
$edgeA numeric matrix with two columns listing the updated ID of rootward and tipward nodes of the remaining edges$edge.lengthA numeric vector. Updated length of remaining edges.$NnodeInteger. Number of remaining internal nodes.$tip.labelCharacter vector. Labels of the branches cut at thefocal_time.If
keep_tip_labels = TRUE, the cut terminal branches are labeled with the tip.label of their unique descendant tip.If
keep_tip_labels = FALSE, the cut terminal branches are labeled with the node ID of their unique descendant tip.
$mapsAn updated list of named numeric vectors. Provides the mapping of trait values along each remaining edge.$mapped.edgeAn updated matrix. Provides the evolutionary time spent across trait values (columns) along the remaining edges (rows).
Additional elements:
$root_ageInteger. Stores the age of the root of the tree.$nodes_ID_dfData.frame with two columns. Provides the conversion from thenew_node_IDto theinitial_node_ID. Each row is a node.$initial_nodes_IDCharacter vector. Provides the initial ID of internal nodes. Used to plot internal node IDs as labels withape::nodelabels().$edges_ID_dfData.frame with two columns. Provides the conversion from thenew_edge_IDto theinitial_edge_ID. Each row is an edge/branch.$initial_edges_IDCharacter vector. Provides the initial ID of edges/branches. Used to plot edge/branch IDs as labels withape::edgelabels().
Details
The mapped phylogenetic tree is cut for a specific time in the past (i.e. the focal_time).
When a branch with a single descendant tip is cut and keep_tip_labels = TRUE,
the leaf left is labeled with the tip.label of the unique descendant tip.
When a branch with a single descendant tip is cut and keep_tip_labels = FALSE,
the leaf left is labeled with the node ID of the unique descendant tip.
In all cases, when a branch with multiple descendant tips (i.e., a clade) is cut, the leaf left is labeled with the node ID of the MRCA of the cut-off clade.
The stochastic trait mapping is updated accordingly by removing mapping associated with the cut off branches.
See also
cut_phylo_for_focal_time() extract_most_likely_trait_values_for_focal_time() extract_all_trait_values_for_focal_time()
For a guided tutorial, see this vignette: vignette("cut_phylogenies", package = "deepSTRAPP")
Examples
# ----- Prepare data ----- #
## Load mammals phylogeny and data from the R package motmot, and implemented in deepSTRAPP
# Data source: Slater, 2013; DOI: 10.1111/2041-210X.12084
data("mammals", package = "deepSTRAPP")
# Obtain mammal tree
mammals_tree <- mammals$mammal.phy
# Convert mass data into categories
mammals_mass <- setNames(object = mammals$mammal.mass$mean,
nm = row.names(mammals$mammal.mass))[mammals_tree$tip.label]
mammals_data <- mammals_mass
mammals_data[seq_along(mammals_data)] <- "small"
mammals_data[mammals_mass > 5] <- "medium"
mammals_data[mammals_mass > 10] <- "large"
table(mammals_data)
#> mammals_data
#> large medium small
#> 36 83 92
# (May take several seconds to run)
# Produce densityMaps using stochastic character mapping based on an equal-rates (ER) Mk model
mammals_cat_data <- prepare_trait_data(tip_data = mammals_data, phylo = mammals_tree,
trait_data_type = "categorical",
evolutionary_models = "ER",
nb_simulations = 100,
return_simmaps = TRUE,
plot_map = FALSE)
#>
#> 2026-10-14 09:38:11.002631 - Fit 1 evolutionary model(s): ER.
#>
#> ------ ER model ------
#>
#> GEIGER-fitted comparative model of discrete data
#> fitted Q matrix:
#> large medium small
#> large -0.006199429 0.003099714 0.003099714
#> medium 0.003099714 -0.006199429 0.003099714
#> small 0.003099714 0.003099714 -0.006199429
#>
#> model summary:
#> log-likelihood = -152.749035
#> AIC = 307.498071
#> AICc = 307.517210
#> free parameters = 1
#>
#> Convergence diagnostics:
#> optimization iterations = 100
#> failed iterations = 0
#> number of iterations with same best fit = 100
#> frequency of best fit = 1.000
#>
#> object summary:
#> 'lik' -- likelihood function
#> 'bnd' -- bounds for likelihood search
#> 'res' -- optimization iteration summary
#> 'opt' -- maximum likelihood parameter estimates
#>
#> 2026-10-14 09:38:12.562326 - Compare model fits.
#>
#> model logL k AIC AICc delta_AICc Akaike_weights rank
#> ER ER -152.749 1 307.4981 307.5172 0 100 1
#> 2026-10-14 09:38:12.563774 - Run simulations for stochastic mapping.
#>
#> make.simmap is sampling character histories conditioned on
#> the transition matrix
#>
#> Q =
#> large medium small
#> large -0.006199429 0.003099714 0.003099714
#> medium 0.003099714 -0.006199429 0.003099714
#> small 0.003099714 0.003099714 -0.006199429
#> (specified by the user);
#> and (mean) root node prior probabilities
#> pi =
#> large medium small
#> 0.3333333 0.3333333 0.3333333
#> Done.
#> 2026-10-14 09:38:24.675632 - Extract ACE as posterior sampling from stochastic mapping.
#> 2026-10-14 09:38:25.084629 - Create densityMaps by summarizing simulations of evolutionary history (simmaps).
#>
#> 2026-10-14 09:38:26.241162 - Posterior probability computed for edge n°100/420
#> 2026-10-14 09:38:27.064068 - Posterior probability computed for edge n°200/420
#> 2026-10-14 09:38:27.907374 - Posterior probability computed for edge n°300/420
#> 2026-10-14 09:38:28.88604 - Posterior probability computed for edge n°400/420
#> 2026-10-14 09:38:29.24301 - Posterior probabilities computed for State = large - n°1/3
#> 2026-10-14 09:38:30.441345 - Posterior probability computed for edge n°100/420
#> 2026-10-14 09:38:31.191823 - Posterior probability computed for edge n°200/420
#> 2026-10-14 09:38:32.157356 - Posterior probability computed for edge n°300/420
#> 2026-10-14 09:38:33.171883 - Posterior probability computed for edge n°400/420
#> 2026-10-14 09:38:33.502907 - Posterior probabilities computed for State = medium - n°2/3
#> 2026-10-14 09:38:34.679853 - Posterior probability computed for edge n°100/420
#> 2026-10-14 09:38:35.473552 - Posterior probability computed for edge n°200/420
#> 2026-10-14 09:38:36.43136 - Posterior probability computed for edge n°300/420
#> 2026-10-14 09:38:37.52781 - Posterior probability computed for edge n°400/420
#> 2026-10-14 09:38:37.845801 - Posterior probabilities computed for State = small - n°3/3
# Extract the simulated evolutionary histories (simmaps)
mammals_simmaps <- mammals_cat_data$simmaps
# Extract simulation n°1
mammals_simmap_1 <- mammals_simmaps[[1]]
# Set focal time to 80Mya
focal_time <- 80
# ----- Example 1: keep_tip_labels = TRUE ----- #
# Cut the simmap to 80 Mya while keeping tip.label
# on terminal branches with a unique descending tip.
updated_simmap_1 <- cut_simmap_for_focal_time(simmap = mammals_simmap_1,
focal_time = focal_time,
keep_tip_labels = TRUE)
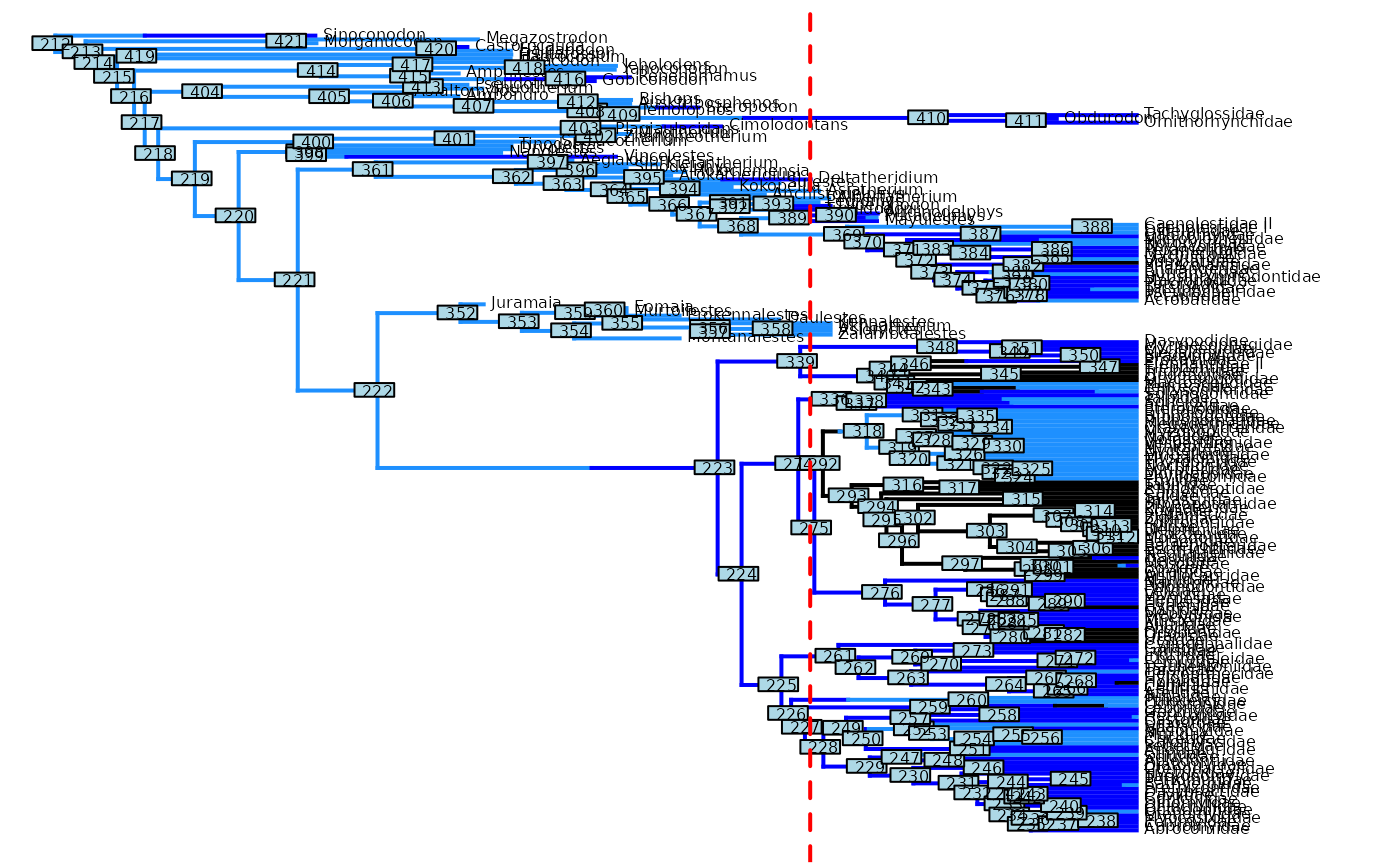
# Plot node labels on initial stochastic map with cut-off
plot(mammals_simmap_1, lwd = 2, fsize = 0.5,
colors = c(large = "black", medium = "blue", small = "dodgerblue"))
ape::nodelabels(cex = 0.5)
abline(v = max(phytools::nodeHeights(mammals_simmap_1)[,2]) - focal_time,
col = "red", lty = 2, lwd = 2)
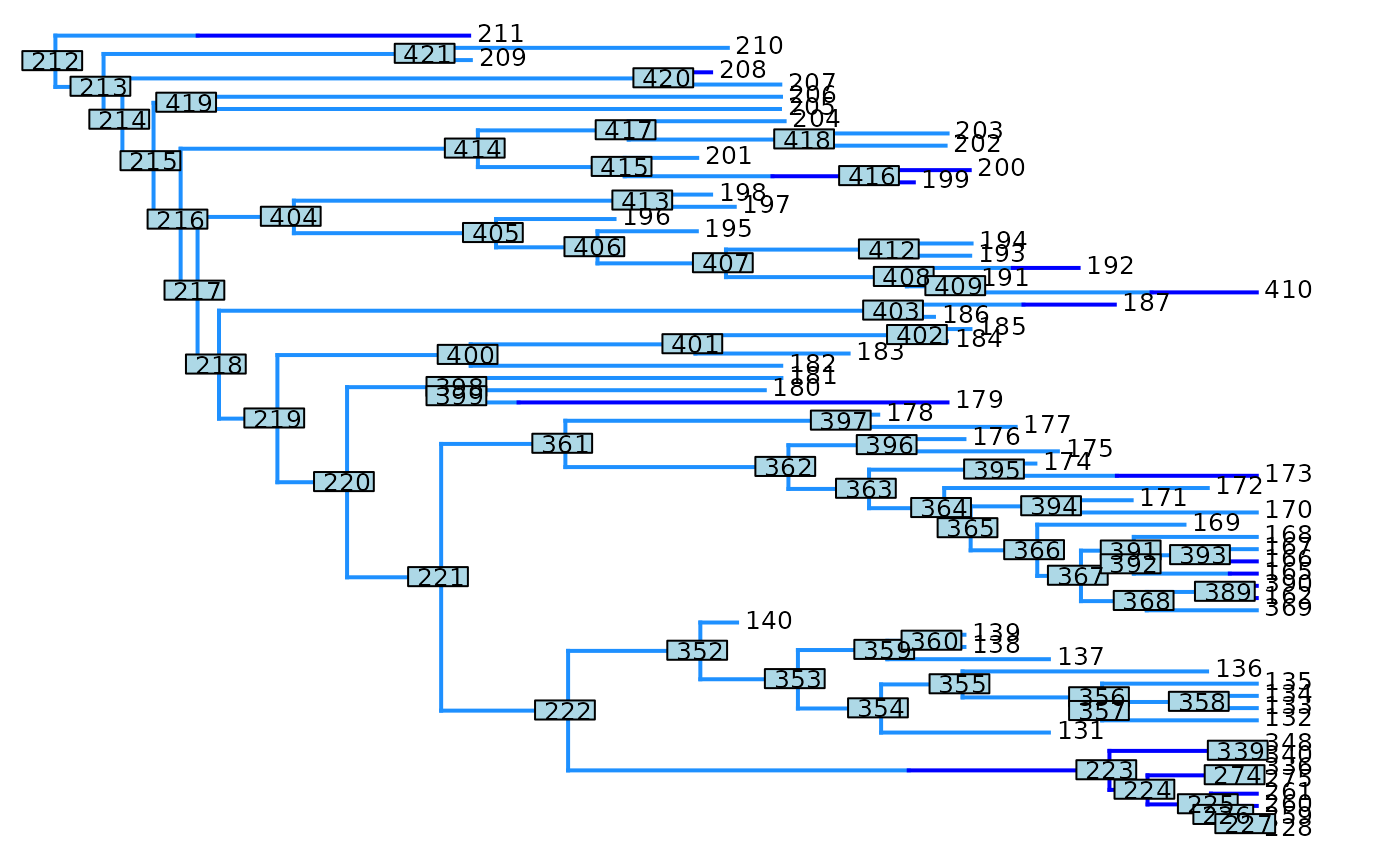
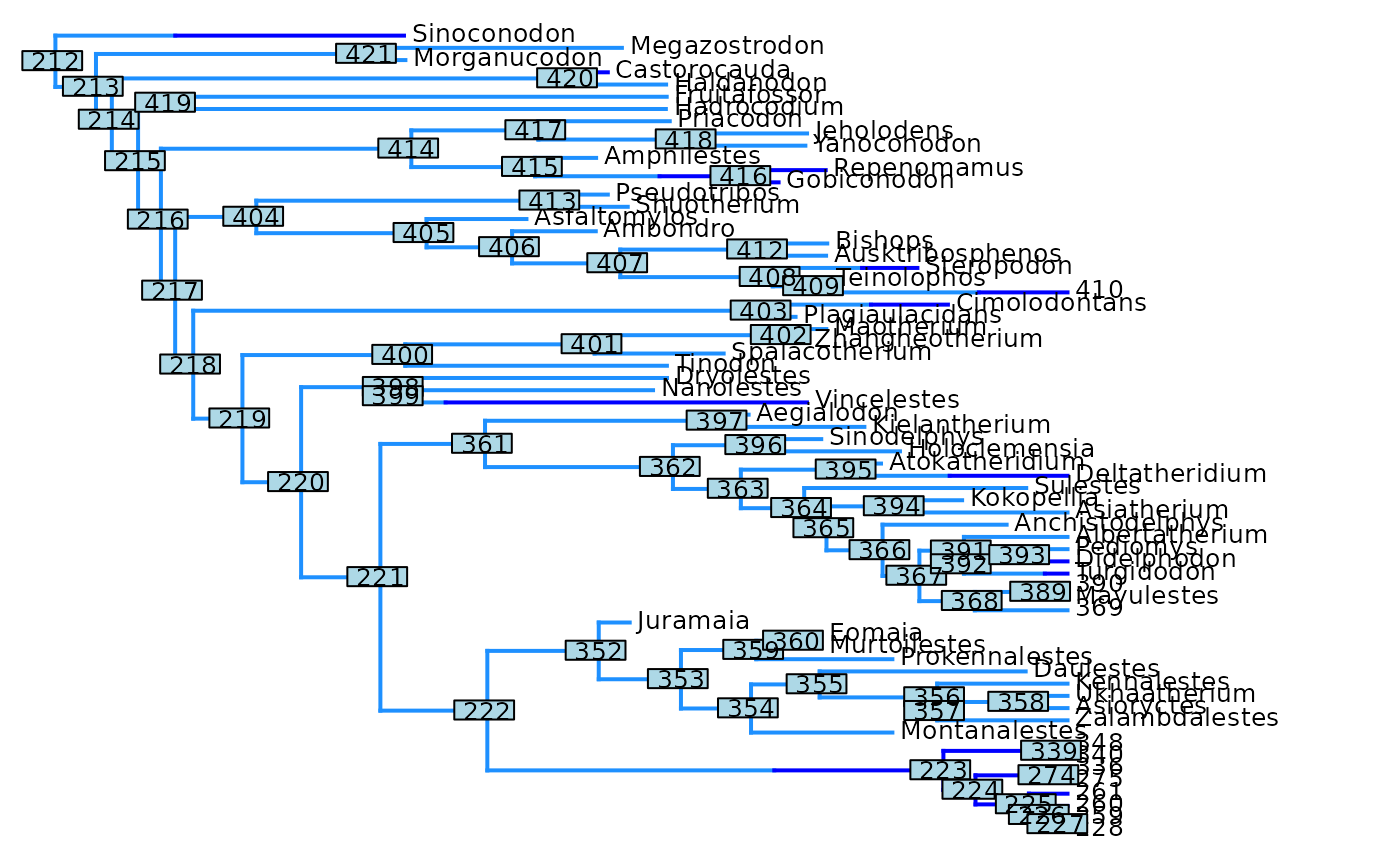
 # Plot initial node labels on cut stochastic map
plot(updated_simmap_1, fsize = 0.8,
colors = c(large = "black", medium = "blue", small = "dodgerblue"))
ape::nodelabels(cex = 0.8, text = updated_simmap_1$initial_nodes_ID)
 # ----- Example 2: keep_tip_labels = FALSE ----- #
# Cut the simmap to 80 Mya while NOT keeping tip.label.
updated_simmap_1 <- cut_simmap_for_focal_time(simmap = mammals_simmap_1,
focal_time = focal_time,
keep_tip_labels = FALSE)
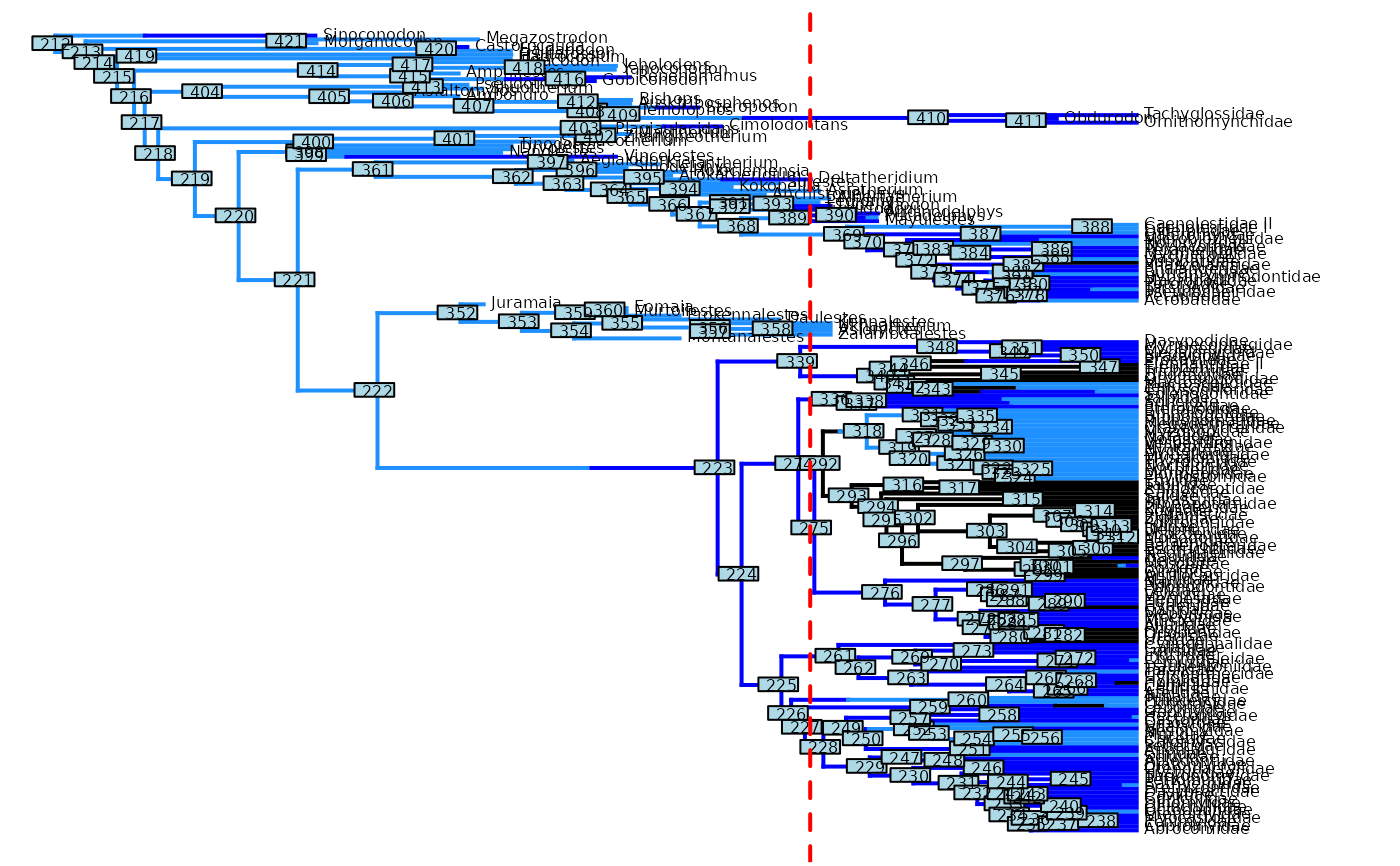
# Plot node labels on initial stochastic map with cut-off
plot(mammals_simmap_1, lwd = 2, fsize = 0.5,
colors = c(large = "black", medium = "blue", small = "dodgerblue"))
ape::nodelabels(cex = 0.5)
abline(v = max(phytools::nodeHeights(mammals_simmap_1)[,2]) - focal_time,
col = "red", lty = 2, lwd = 2)
 # Plot initial node labels on cut stochastic map
plot(updated_simmap_1, fsize = 0.8,
colors = c(large = "black", medium = "blue", small = "dodgerblue"))
ape::nodelabels(cex = 0.8, text = updated_simmap_1$initial_nodes_ID)